Gedatolisib (PKI-587) Gedatolisib from Selleck Dual PI3K/mTOR Inhibition Reverses Cetuximab Resistance
Copy code to add this image to your page
Head and neck squamous cell carcinoma (HNSCC) frequently displays overexpression of the epidermal growth factor receptor (EGFR). Monoclonal antibodies targeting EGFR, such as cetuximab, initially provided a targeted therapeutic option for patients with advanced disease. However, persistent intrinsic or acquired resistance severely limits long-term clinical efficacy. Biochemical investigations demonstrate that resistance often emerges from compensatory activation of downstream survival networks that bypass EGFR blockade. Chief among these bypass pathways is the phosphoinositide 3-kinase (PI3K) / Akt / mammalian target of rapamycin (mTOR) axis. When PI3K signaling remains constitutively active, tumor cells sustain survival signaling despite upstream receptor neutralization. Consequently, single-agent therapy against EGFR proves insufficient to control refractory tumor growth.
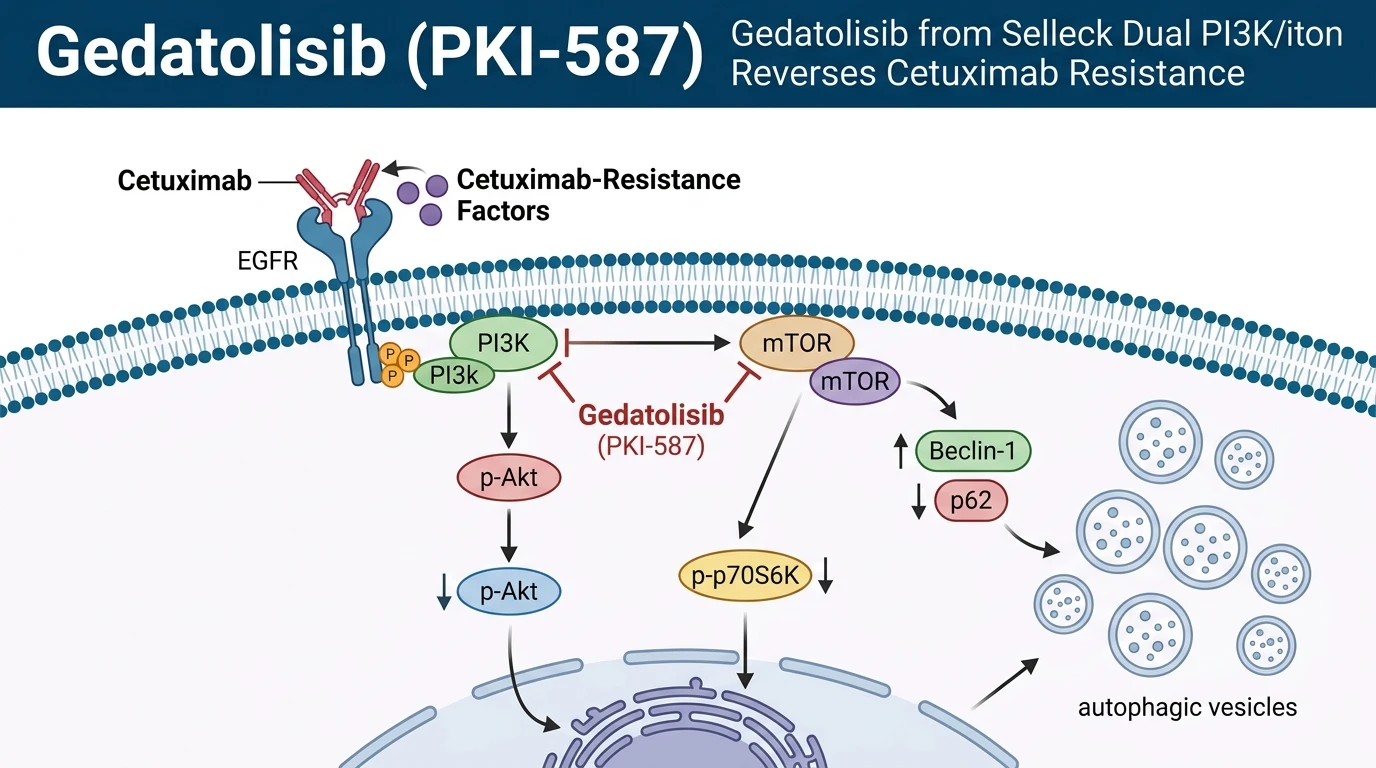
To counter this signaling bypass, targeting downstream nodes directly offers a rational therapeutic strategy. Gedatolisib (PKI-587), a potent dual inhibitor of both class I PI3K isoforms and mTOR complexes (mTORC1 and mTORC2), was developed to prevent selective feedback activation that often compromises single-node inhibitors. High-throughput compound screening identified Gedatolisib from Selleck as an effective agent to suppress concurrent PI3K and mTOR signaling events. By concurrently occupying the ATP-binding pockets of PI3K catalytic subunits and the mTOR kinase domain, Gedatolisib (PKI-587) prevents the phosphorylation of Akt at both Ser473 and Thr308 residues, while simultaneously blocking p70S6K activation at Thr389. In experimental HNSCC models, this dual blockade disrupts protein translation and cellular proliferation.
Mechanistic evaluation reveals distinct death mechanisms elicited by dual PI3K/mTOR targeted therapy depending on the baseline cetuximab sensitivity of the target cells. In cetuximab-sensitive HNSCC cell lines, combined treatment with cetuximab and Gedatolisib (PKI-587) induces canonical caspase-dependent apoptosis. This phenotype is characterized by cleavage of poly(ADP-ribose) polymerase (PARP), activation of caspase-3 and caspase-9, and rapid annexin V externalization. Conversely, in cetuximab-resistant HNSCC cell lines, dual inhibition prompts a shift in death pathways. Rather than executing primary apoptosis, resistant cells undergo extensive autophagic remodeling. Treatment elevates expression of the pro-autophagic protein Beclin-1 while promoting the degradation and depletion of p62/SQSTM1. This alteration in autophagic flux serves as a primary driver of cell death in context-dependent models where apoptotic execution machinery is blunted or downstream anti-apoptotic proteins are overexpressed.
In vivo proof-of-concept studies utilizing Kyse30 xenograft murine models confirm the physiological relevance of these cellular observations. Mice bearing cetuximab-resistant human carcinoma xenografts received monotherapy or combination regimens under controlled experimental parameters. Monotherapy with either cetuximab or dual PI3K/mTOR inhibition produced modest growth deceleration. However, combined administration of Gedatolisib (PKI-587) with cetuximab generated marked tumor regression and significantly extended overall survival compared to vehicle controls. Biochemical profiling of harvested xenograft tissues verified sustained suppression of p-Akt and p-p70S6K throughout the dosing schedule. These preclinical data confirm that co-targeting PI3K and mTOR restores tumor vulnerability to cetuximab in refractory solid cancers.
Pharmacological Characterization and Resistance Mitigation Strategies
Copy code to add this image to your page
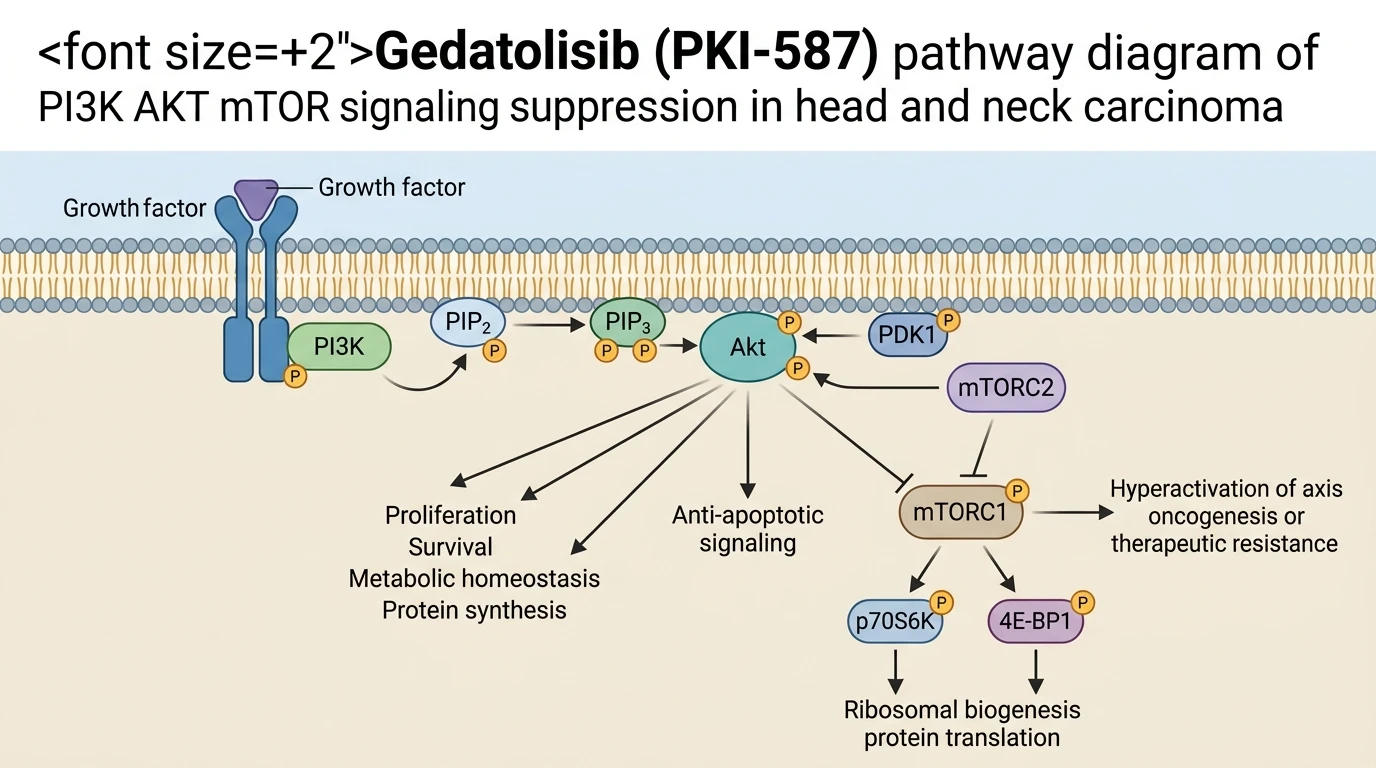
Gedatolisib (PKI-587) functions as a dual ATP-competitive inhibitor of class I PI3K catalytic isoforms and mTOR complexes (mTORC1/mTORC2). By simultaneously blocking both kinase nodes, it eliminates compensatory Akt phosphorylation at Ser473/Thr308 and suppresses p70S6K activity at Thr389. In EGFR-inhibitor sensitive HNSCC cells, this blockade activates caspase-dependent apoptotic pathways. In cetuximab-resistant cells, it triggers autophagy characterized by elevated Beclin-1 and depleted p62 levels. This dual mechanism bypasses upstream EGFR resistance and suppresses tumor progression in vivo.
Dual PI3K and mTOR Suppression Attenuates Akt Phosphorylation
Targeting single signaling nodes within the PI3K/Akt/mTOR pathway frequently induces loss of negative feedback loops. This loss leads to compensatory Akt reactivation via mTORC2 or receptor tyrosine kinase upregulation. Gedatolisib from Selleck circumvents this compensatory mechanism through simultaneous inhibition of class I PI3K isoforms and mTORC1/mTORC2 catalytic centers. In vitro biochemical assays performed on cetuximab-resistant HNSCC cell lines demonstrate rapid, concentration-dependent suppression of Akt phosphorylation at Ser473 and Thr308. Downstream effector activity ceased within two hours of drug exposure. Phosphorylation of p70S6K at Thr389 was completely abated at nanomolar concentrations. Proliferation assays confirmed a marked reduction in colony formation following treatment. The dual-binding kinetic profile ensures that signal propagation downstream of mutated or amplified PI3K catalytic subunits remains completely blocked, neutralizing the primary driver of acquired cetuximab resistance.
Autophagic Remodeling Drives Re-Sensitization in Resistance Models
Cetuximab-resistant cell lines exhibit altered programmed cell death pathways upon exposure to dual PI3K/mTOR blockade. Cellular viability dropped sharply in a dose-dependent manner. While cetuximab-sensitive cells engage canonical caspase cascades, resistant models display pronounced autophagic flux. Immunoblot analysis revealed significant upregulation of Beclin-1 expression following exposure to Gedatolisib (PKI-587). Simultaneously, p62/SQSTM1 protein levels decreased substantially, indicating uninterrupted lysosomal degradation of autophagosomes. Pharmacological inhibition of early-stage autophagy attenuated drug-induced lethality in these resistant contexts. These observations demonstrate that autophagic execution acts as a primary cytotoxic mechanism when apoptotic induction is compromised. Re-sensitization to cetuximab relies directly on this autophagic shift under tested in vitro assay conditions.
Kyse30 Xenograft Regressions Confirm In Vivo Antitumor Efficacy
In vivo evaluation was conducted using immunocompromised mice bearing Kyse30 human carcinoma xenografts. Treatment cohorts were assigned to evaluate vehicle, cetuximab monotherapy, dual PI3K/mTOR inhibitor monotherapy, or combined therapy. Monotherapy arms demonstrated transient tumor growth retardation but failed to prevent disease progression. Combined treatment with Gedatolisib (PKI-587) and cetuximab resulted in significant tumor volume reduction over a 28-day administration protocol. Animals receiving combination therapy exhibited statistically significant survival extension compared to monotherapy controls. Immunohistochemical staining of excised tumor tissues confirmed robust down-regulation of p-Akt and p-p70S6K. Systemic toxicity remained low, with animal body weights remaining stable throughout the study period. These preclinical findings establish dual PI3K/mTOR targeting as a viable therapeutic strategy to overcome EGFR targeted therapy resistance.