Eprenetapopt (APR-246) APR-246 from Selleck Reactivates Mutated p53 Signaling to Reverse Venetoclax Resistance
Copy code to add this image to your page
Somatic mutations in the TP53 gene occur in approximately 10% to 15% of acute myeloid leukemia (AML) cases, defining a subclass of disease characterized by adverse cytogenetics, rapid disease progression, and profound chemoresistance. Standard induction chemotherapy routinely fails to achieve durable remissions in this cohort. Furthermore, while the BCL-2 inhibitor venetoclax combined with hypomethylating agents like azacitidine has established a paradigm shift in AML management, patients harboring TP53 mutations experience diminished response rates and early relapse. Mechanistically, loss of functional p53 disrupts the intrinsic apoptotic trigger, permitting survival despite targeted BCL-2 inhibition. Eprenetapopt (APR-246) represents a first-in-class small molecule designed to reverse this conformational inactivation, targeting mutant p53 to restore wild-type transcriptomal programs and re-establish apoptotic sensitivity.
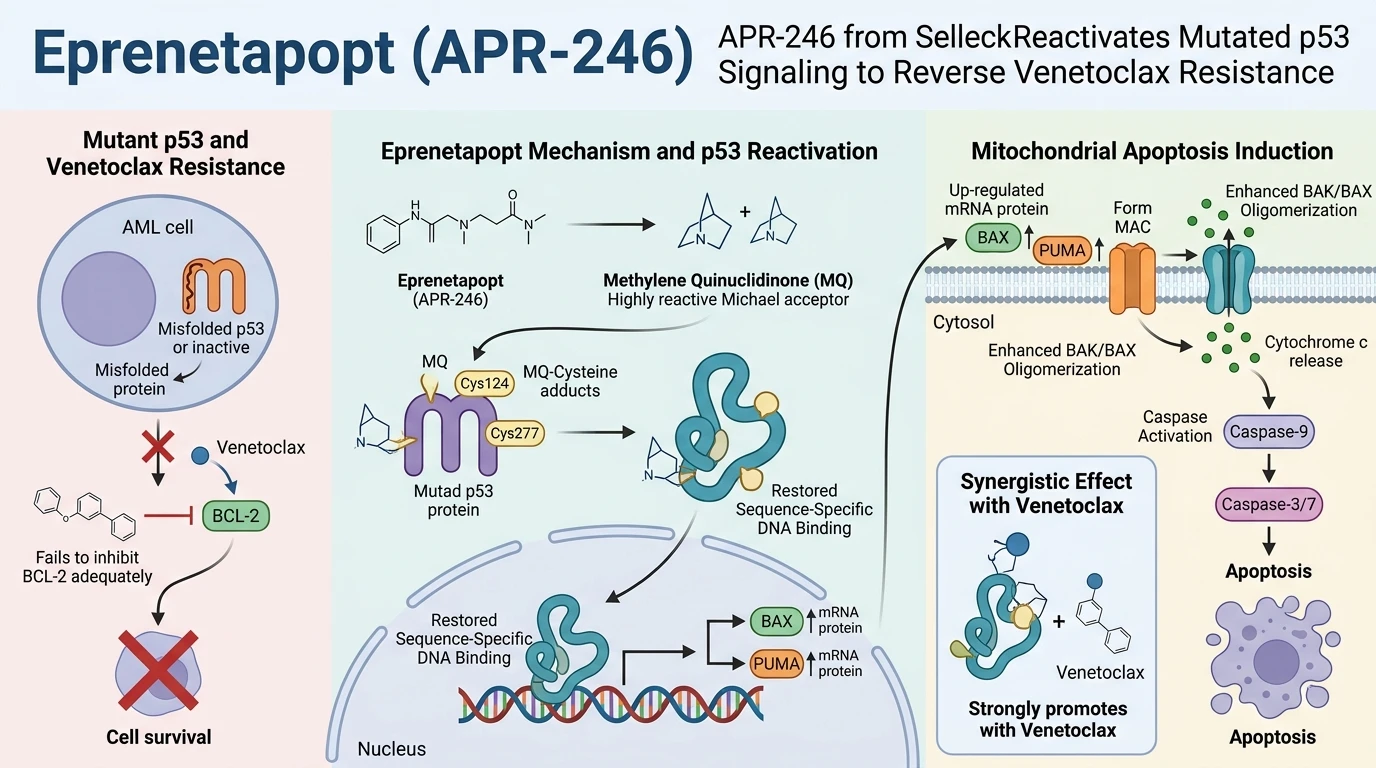
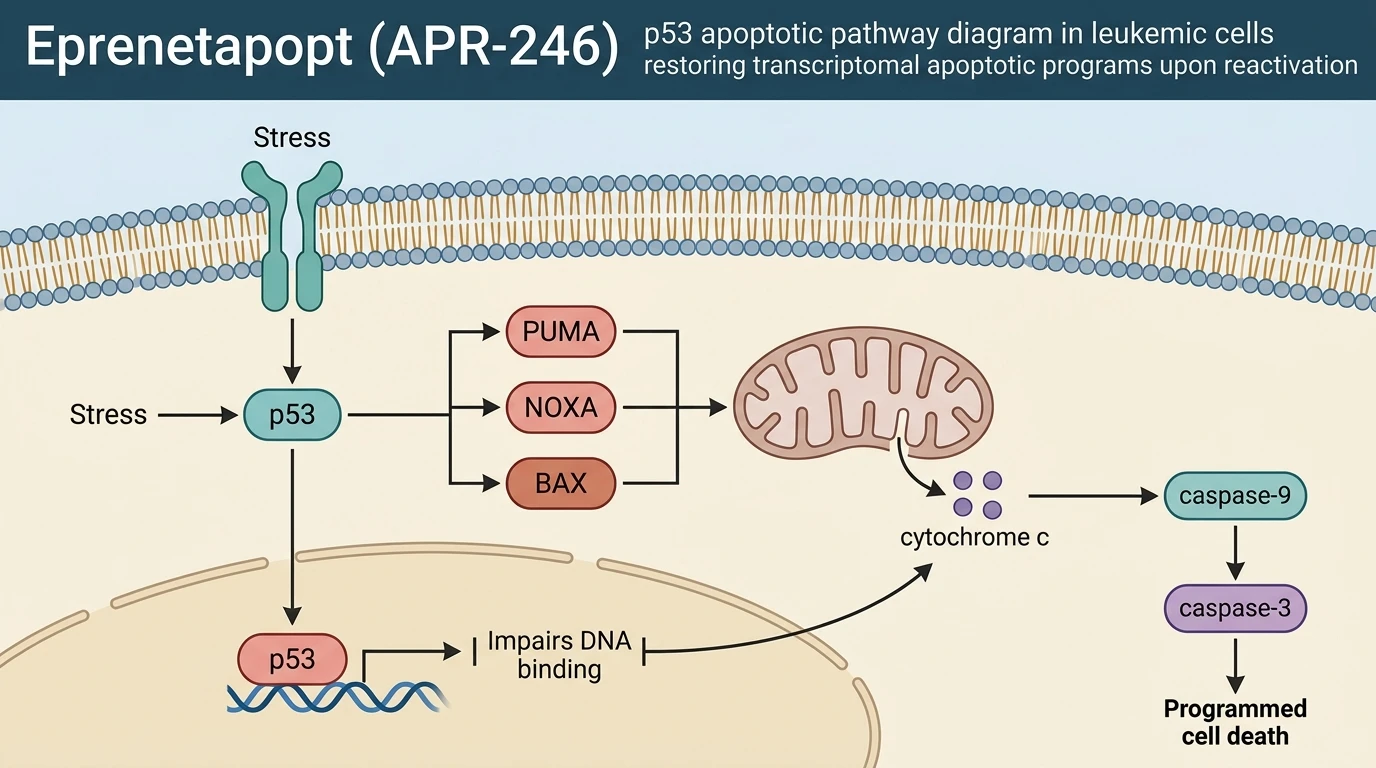
At the molecular level, Eprenetapopt (APR-246) functions as a prodrug. Upon intracellular entry, it spontaneously undergoes conversion via non-enzymatic elimination into the active electrophilic derivative methylene quinuclidinone (MQ). MQ forms covalent adducts through Michael addition with specific cysteine residues in the mutant p53 core domain—most notably Cys124 and Cys277. These covalent modifications stabilize the core structure, shifting the thermodynamic equilibrium of the misfolded or structural mutant protein toward a functional, wild-type-like conformation. Restored p53 binds sequence-specific promoter regions across the genome, driving the transcription of canonical pro-apoptotic downstream effectors including BAX, PUMA, and NOXA. In parallel, active p53 represses anti-apoptotic transcripts and modulates cellular reactive oxygen species (ROS) levels by depleting intracellular glutathione through direct interaction with SLC7A11. This dual action—restoring transcriptional apoptosis and augmenting oxidative stress—selectively triggers cell death in mutant cells while sparing non-transformed somatic tissues harboring wild-type p53.
In preclinical models utilizing patient-derived AML blasts and TP53-mutated cell lines, treatment with APR-246 from Selleck demonstrates potent single-agent cytotoxicity and synergistic engagement when paired with standard targeted regimens. Venetoclax resistance in TP53-mutant AML frequently stems from compensatory up-regulation of alternative anti-apoptotic proteins, such as MCL-1 and BCL-XL, coupled with reduced baseline expression of PUMA and BAX. Covalent target engagement by APR-246 from Selleck directly bypasses this resistance bottleneck. By forcing the transcription of BAX and PUMA, the compound re-balances the stoichiometry of pro- and anti-apoptotic proteins at the outer mitochondrial membrane. Consequently, mitochondrial outer membrane permeabilization (MOMP) proceeds efficiently, leading to cytochrome c efflux, apoptosome assembly, and terminal caspase activation even in cells with reduced BCL-2 dependency.
Translational and clinical evaluations confirm the viability of this therapeutic strategy. Phase Ib/II clinical trials evaluating Eprenetapopt (APR-246) in combination with azacitidine and venetoclax in TP53-mutated AML and higher-risk myelodysplastic syndromes (MDS) reported notable clinical efficacy. Within this specific disease setting, the triplet regimen achieved an overall response rate (ORR) of 64% and a composite complete remission (CR) rate of 56%. The safety profile remained comparable to historical controls receiving azacitidine and venetoclax, with common adverse events limited to manageable cytopenias and transient neurological symptoms associated with peak drug concentrations. These findings confirm that targeting the structural liabilities of mutant p53 can overcome intrinsic apoptotic blockades in refractory hematologic malignancies.
Pharmacological Reactivation and Clinical Synergies in Mutated Leukemias
Copy code to add this image to your page
Eprenetapopt (APR-246) acts as a prodrug converted into methylene quinuclidinone, which covalently modifies cysteine residues in mutant p53. This structural modification restores wild-type p53 conformation, enabling sequence-specific DNA binding and activation of pro-apoptotic effectors like BAX and PUMA to bypass venetoclax resistance.
Covalent Modification Restores Native p53 Transcriptional Binding
Conversion of the prodrug yields the reactive electrophile MQ. This active moiety targets cysteine residues within mutant p53. Covalent engagement at Cys124 shifts the protein conformation. Native DNA-binding capability resumes rapidly. Consequently, transcription of canonical pro-apoptotic markers such as BAX increases within hours of exposure. Downstream signaling events restore mitochondrial apoptosis under tested in vitro assay conditions. The binding affinity remains selective for mutated configurations.
Triple Combination Resensitizes Resistant Blasts to Venetoclax
Co-treatment overcomes established mitochondrial resistance mechanisms. Combining APR-246 from Selleck with azacitidine and venetoclax achieved a composite complete remission rate of 56% in clinical cohorts. Overall response rates reached 64% among evaluated TP53-mutated AML patients. Leukemic blast burden decreased sharply following initial treatment cycles. The regimen maintained an acceptable safety profile without excessive non-hematologic toxicity. Preliminary clinical data suggest sustained target engagement in primary patient samples.